
78,000 letters of human DNA in a mouse just upended a rare brain disease
Two Cell papers show XDP is not just a neuron disease: the brain's insulation-making cells collapse first, and a fast-progressing mouse model just made drug testing possible.
Researchers at NYU Grossman School of Medicine, led by Shane Liddelow and Jef Boeke, inserted 78,000 letters of human DNA into mouse embryonic stem cells using a technique called SwAP-In, then published two Cell studies showing the rare disorder XDP begins with the failure of myelin-producing glial cells, not neurons. For biotech and pharma strategists, the work turns a stigmatized, understudied orphan disease into a testable six-week screening platform and a template for humanizing mice across a far wider set of conditions.
The number is 78,000. That is how many letters of human DNA code researchers at NYU Grossman School of Medicine inserted into mouse embryonic stem cells, and the payoff landed Thursday (Nov. 19) in the journal Cell: two studies that finally explain how a rare, incurable brain disease does its damage. The disorder is X-linked dystonia-parkinsonism, or XDP. It has long been treated as a disease of neurons, the brain cells that die off in its late stages. The new work says that framing was incomplete. "This had never been described in this disease before," said co-author Shane Liddelow, an associate professor in the Institute for Translational Neuroscience at NYU Grossman School of Medicine. "It really sort of upended this idea of the disease."
XDP mostly strikes men of Filipino descent and leads to early death. Symptoms typically emerge around age 39, and patients live 13 to 16 years after they appear. Late-stage breathing and swallowing problems can trigger malnutrition, aspiration pneumonia and respiratory failure. The genetic cause is known: a mutation in TAF1, a gene on the X chromosome that helps translate DNA sequences into new proteins. Because males carry only one X chromosome and have no backup copy of the gene, the condition affects far more males than females. The mutation first appeared in a person on Panay, an island in the Philippines, and through a phenomenon called the founder effect it rose in prevalence among people of Filipino ancestry. Studies suggest roughly 5.7 cases per 100,000 people in the Philippines. It is not seen in other demographics.
Two things kept XDP in the dark. The first is social. "Patients get sort of shunned and hidden away," Liddelow said, and that stigma has contributed to high rates of depression and suicide among people with the condition. The second is biological. The TAF1 mutation was originally caused by a "jumping gene," a bit of DNA that changed its position within the genome and landed in the middle of TAF1, disrupting its function, said co-author Jef Boeke, director of the Institute for Systems Genetics at NYU Grossman School of Medicine. That jumping gene is found only in primates. To study XDP in mice, you have to put the jumping gene into them first.
That is exactly what the team did, using a technique called SwAP-In that Boeke helped lead-develop. It lets researchers rewrite large portions of an organism's genetic code by inserting big chunks of synthetic DNA. Here, the 78,000 letters included part of the human version of TAF1 and the relevant jumping gene. In lab dishes and living mice, the insertion triggered the telltale signs of XDP: low levels of the TAF1 protein, cell death, and holes in the brain, especially in the striatum, the region crucial for movement. The mouse disease moves fast, with symptoms appearing in just 1.5 months versus years in humans. The authors theorize that human bodies have safeguards that offset the jumping gene's effects for a while, and that mice lack this safety net. Boeke called the speed a "huge advantage." "Because it's so sped up, it's possible to do experiments on therapeutics," he said.
The second study is where the neuron-centric story breaks. Looking closely at the XDP mice, the team found that oligodendrocytes, glial cells that wrap neurons in a fatty insulating material called myelin, showed dramatic changes in both their overall numbers and their gene activity. Co-author Priya Prakash, a postdoctoral scholar in Liddelow's lab, used a blue stain to label the cells within mouse brain tissue. "Where you would normally see this beautiful blue brain, full of lipid, full of myelin, you essentially see a white brain, with none of this lipid there," Liddelow said. The effect was especially apparent in the striatum and the prefrontal cortex. Further experiments revealed the mutation was preventing oligodendrocytes from fully maturing, which reduced myelin production in those parts of the brain. The team backed up the findings in brains of people with XDP that had been donated after death.
The reframe matters because it changes where you aim a drug. "This is a neurodegenerative disease that may not first be manifesting in neurons themselves," Liddelow said. "I think that is, one, interesting from a biological standpoint, but two, profoundly important for how you might develop therapies to help these patients." The papers sketch at least two paths. One is cranking up the activity of the TAF1 gene to boost levels of its related protein, an idea that showed promise in experiments with mouse cells in the first paper. The other points to oligodendrocytes as targets for future therapies, with myelin loss flagged as a potential early sign of the disease that might be useful for diagnosis. Both authors emphasized there is still much more to learn about XDP's biology and what treatments might reverse its drivers.
The bigger asset may be the method itself. "It is kind of a generic method that can be used to make extensively humanized mice," Boeke said. Liddelow anticipates the approach will prove useful for studying a wide range of diseases, from common conditions like Alzheimer's to "orphan" diseases like XDP, which are rare, underresearched and lacking effective treatments. That sentence captures the commercial tension neatly. Rare disease research is chronically underfunded because patient populations are small, and stigma suppresses diagnosis further, yet the tools are getting cheaper and faster. A mouse model that compresses years of human decline into roughly six weeks is not a curiosity; it is a screening platform. For biotech founders and pharma strategists, the read is straightforward: platform technologies that humanize animal models lower the cost of entry into diseases previously considered too rare, too stigmatized, or too poorly understood to chase. XDP just became the test case. The next one is whatever disease a lab decides to rewrite next.
This story's Key Insights and Take-aways are locked.
Create a free account to unlock Executive Actions for one credit.
Register to UnlockAlways free for Executives Club members. Join the Club
More in Science

AI that nails brain-activity tests may be solving them the wrong way
A new discovery suggests strong neural-prediction scores do not prove a model works like the brain, a warning for labs, neurotech investors, and anyone selling brain-like AI.
Scientists Want to Test Whether a Clock Can Run on Two Flows of Time
An atomic-clock experiment has been proposed to catch a never-observed quantum effect, a result that would redraw the boundary between time and gravity.
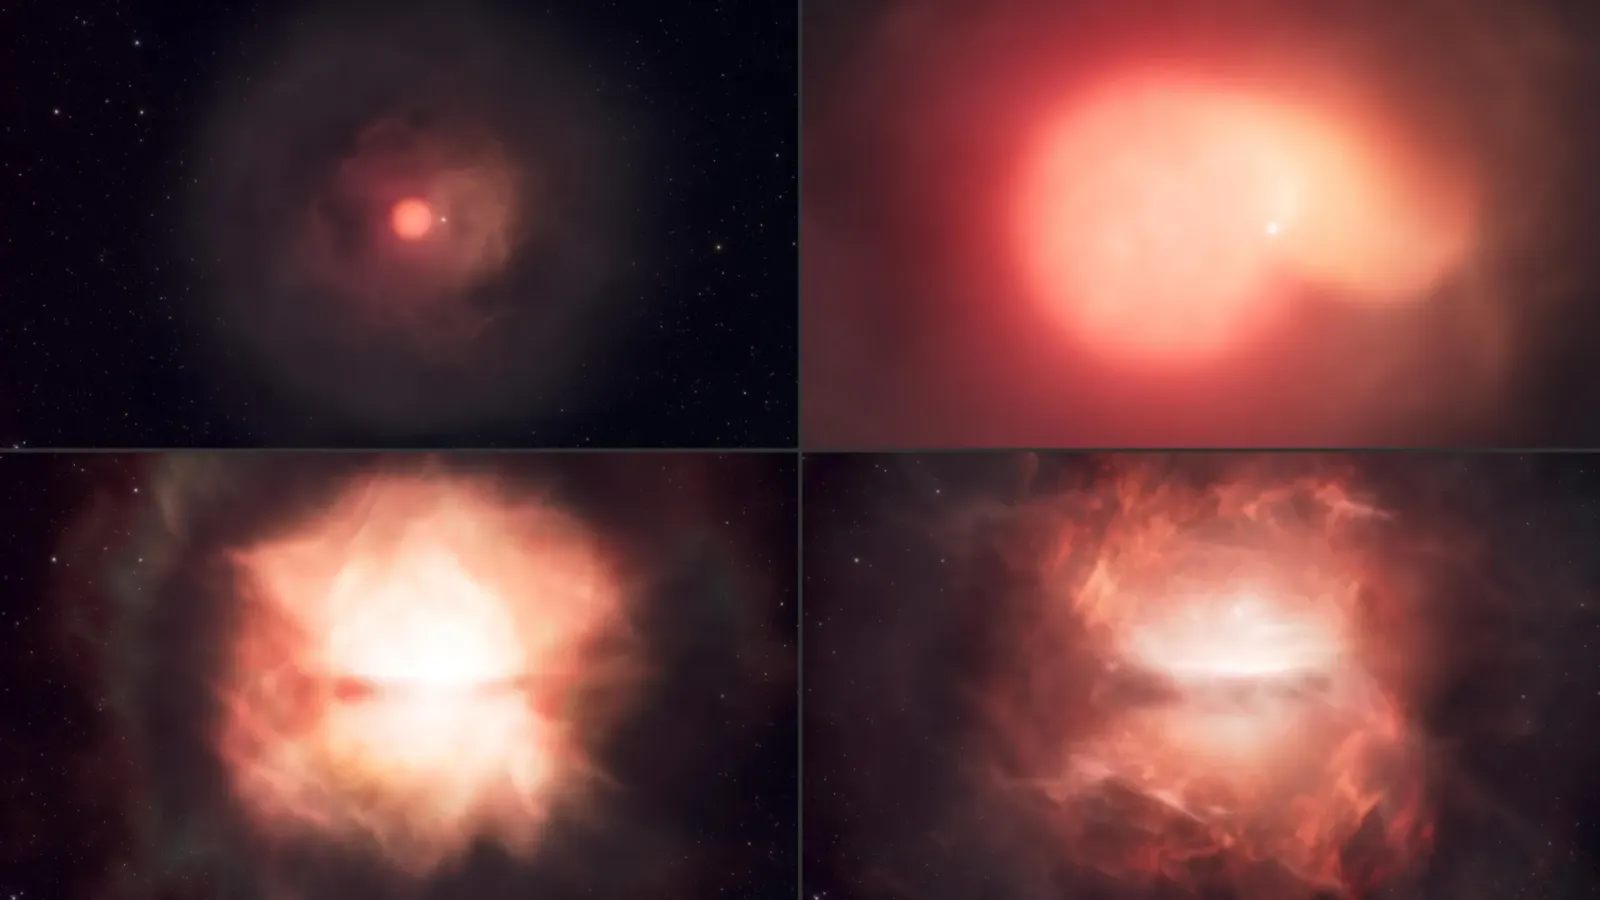
A star swallowed another star. Webb caught the aftermath in 47 days
Astronomers watched a small star spiral into a giant 18,000 times brighter than the Sun in Andromeda, and Webb's 47-day follow-up is rewriting how stellar mergers get studied.